
核磁共振氢谱和碳谱中各溶剂的化学位移
核磁共振碳谱和氢谱中 化学位移产生的原因

核磁共振(NMR)技术是一种应用广泛的谱学技术,常用于分析有机物和生物分子的结构和性质。
在核磁共振谱中,化学位移是一个重要的参数,它与化合物中原子核周围的电子环境有关。
化学位移在碳谱和氢谱中都是十分常见的,在本文中,我们将探讨化学位移在核磁共振碳谱和氢谱中产生的原因。
1. 基本概念在核磁共振谱中,化学位移是指核磁共振信号的频率与参考物质(通常是三氯化甲烷或二甲基硅烷)信号频率之差。
化学位移通常用ppm (parts per million)表示,它是一个相对值,可以用来比较不同化合物中原子核的化学环境差异。
2. 碳谱中化学位移的影响因素碳谱中的化学位移受到多种因素的影响,其中主要包括化学环境、电子效应和磁场效应。
- 化学环境:不同化学环境下的碳原子核受到不同的化学位移影响。
芳香环上的碳原子与脂肪链上的碳原子所受的化学环境不同,因此它们的化学位移也会有所差异。
- 电子效应:分子中的电子密度分布会影响到周围原子核的化学位移。
含有电子丰富基团的碳原子通常会表现出较低的化学位移,而含有电子贫瘠基团的碳原子则会表现出较高的化学位移。
- 磁场效应:外加磁场对原子核周围的电子运动轨迹会产生影响,从而影响原子核的化学位移。
这种效应在核磁共振谱分析中是不可忽视的。
3. 氢谱中化学位移的影响因素类似于碳谱,氢谱中的化学位移也受到化学环境、电子效应和磁场效应的影响。
- 化学环境:不同化学环境下的氢原子核受到不同的化学位移影响。
α-位置上的氢原子与β-位置上的氢原子所受的化学环境不同,因此它们的化学位移也会有所差异。
- 电子效应:分子中的电子密度分布会影响到周围原子核的化学位移。
对甲苯中的甲基氢和对位氢受到的电子效应不同,因此它们的化学位移也会有所差异。
- 磁场效应:外加磁场对原子核周围的电子运动轨迹会产生影响,从而影响原子核的化学位移。
这种效应在氢谱分析中同样需要考虑。
4. 结语化学位移在核磁共振碳谱和氢谱中的产生是一个复杂而又精密的过程,受到多种因素的影响。
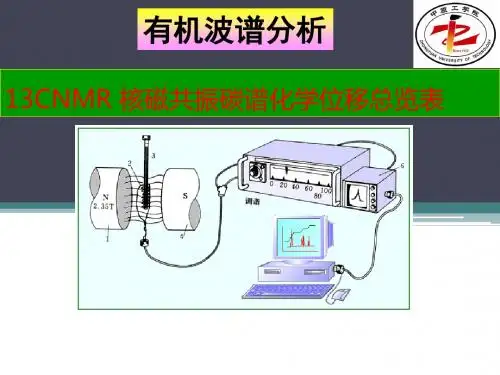
13CNMR 核磁共振碳谱化学位移总览表==
由于15N的天然丰度很小,只有14N的0.37%,因此13C与15N 直接相连的概率很低,偶合常数也很小。一般13C与15N的偶合常数 在1-15Hz。
4.5 核磁共振碳谱解析及应用 4.5.1 核磁共振碳谱谱图解析程序
13C NMR谱的解析并没有一个成熟、统一的程序,应该根 据具体情况,结合其他物理方法和化学方法测定的数据,综合 分析才能得到正确的结论。
by the broadband decoupler
34
4
3 21
56
1
2
5 6
谱图去偶作用对比
6.INEPT谱和DEPT谱
常规的13C NMR谱是指宽带质子去偶谱。在去偶的条件 下,失去了全部C-H偶合的信息,质子偶合引起的多重谱线合 并,每种碳原子只给出一条谱线。虽然用偏共振去偶技术可以 分辨CH3、CH2、CH及季C的归属,但由于偏共振去偶谱中偶合常 数分布不均匀,多重谱线变形和重叠,在复杂分子的研究中仍 然受到限制。随着现代脉冲技术的发展,产生了一些新的能确 定碳原子级数的新方法,如J调制法、APT法、INEPT法和DEPT 法等,其中INEPT法和DEPT法已被广泛应用。
(4)分析偏共振去偶谱和DEPT谱,了解与各种不同化学环境的碳直 接相连的质子数,确定分子中有多少个CH3、CH2、CH和季碳及其可 能的连接方式。比较各基团含H总数和分子式中H的数目,判断是否 存在一OH、一NH2、一C(X)H、一NH一等含活泼氢的基团。 (5)如果样品中不含F、P等原子,宽带质子去偶谱图中的每一条谱 线对应于一种化学环境的碳,对比偏共振去偶谱,全部偶合作用产 生的峰的裂分应全部去除。如果还有谱线的裂分不能去除,应考虑 分子中是否含F或P等元素。 (6)从分子式和可能的结构单元,推出可能的结构式。利用化学位 移规律和经验计算式,估算各碳的化学位移,与实测值比较。 (7)综合考虑1H NMR、IR、MS和UV的分析结果,必要时进行其他的双 共振技术及τ1测定,排除不合理者,得到正确的结构式。
CNMR核磁共振碳谱化学位移总览表医学知识课件
4.3.2 影响 C化学位移的因素 13 文档仅供参考,不能作为科学依据,请勿模仿;如有不当之处,请联系本人改正。
1.碳杂化轨道
碳原子的杂化轨道状态(sp3、sp2、sp)很大程度上决定13C 化学位移。sp3杂化碳的共振信号在高场,sp2杂化碳的共振信号 在低场,sp杂化碳的共振信号介于前二者之间。以TMS为标准, 对于烃类化合物来说,sp3碳的δ值范围在0-60ppm;sp2杂化碳的 δ值范围在100-150ppm,sp杂化碳的δ值范围在60-95ppm。
20多年来核磁共振技术取得巨大发展目前13cnmr已广泛应用于有机化合物的分子结构测定反应机理研究异构体判别生物大分子研究等方面成为化学生物化学药物化学及其他相关领域的科学研究和生产部门不可缺少的分析测试手段对有关学科的发展起了极大的促进作用
4.1 核磁共振碳谱的特点 文档仅供参考,不能作为科学依据,请勿模仿;如有不当之处,请联系本人改正。 13C核磁共振谱的信号是1957年由P. C. Lauterbur首先观察到
3.炔烃的化学位移值
炔基碳为sp杂化,其化学位移介于sp3与sp2杂化碳之间,为 67-92ppm。
4.芳环碳和杂芳环碳的δ 值 文档仅供参考,不能作为科C学依据,请勿模仿;如有不当之处,请联系本人改正。
芳环碳的化学位移值一般在120-160ppm范围内,峰往往出现
在较低场,这点与脂肪族季碳峰在较低场是类似的。
3
4
3 21
56
4
1 2
5 6
文档仅供参考,不能作为科学依据,请勿模仿;如有不当之处,请联系本人改正。
13C NMR spectrum with the protons
氘代dmso化学位移
氘代dmso化学位移
氘代DMSO(dimethylsulfoxide)是一种有机化合物,广泛应用于化学、生物学和材料科学等领域。
氘代DMSO的主要特点是其中一个氧原子被氘原子替换,这使得它在实验研究中具有独特的优势。
氘代DMSO在化学位移中的应用非常广泛。
由于氘原子与氢原子的相互作用,氘代DMSO可以作为一种氘代溶剂,用于研究溶剂效应、氢键和分子间相互作用等。
在核磁共振氢谱(1H-NMR)和核磁共振碳谱(13C-NMR)等实验中,氘代DMSO可以作为溶剂或添加剂,提高样品的溶解度,从而获得更好的信号。
氘代DMSO的优势在于其对许多化合物的溶解度较高,能够显著提高谱图质量。
此外,氘代DMSO可以减少氢氘交换反应,降低信号干扰。
然而,氘代DMSO也存在局限性,如对某些化合物的不稳定性、可能导致样品降解等问题。
因此,在实际应用中,需要根据具体研究目标和实验条件选择合适的氘代溶剂。
在实验中使用氘代DMSO时,应注意以下几点:
1.严格遵循实验规程,确保实验安全。
氘代DMSO具有一定的毒性,应避免直接接触皮肤和眼睛,并在通风良好的环境下操作。
2.选择合适的氘代DMSO浓度。
过高或过低的浓度都可能影响实验结果,因此需要根据具体实验需求调整氘代DMSO的用量。
3.注意氘代DMSO的保存条件。
氘代DMSO应在干燥、避光的环境中储存,并注意密封防潮。
4.考虑到氘代DMSO可能与某些化合物发生相互作用,实验前应对目标化合物进行充分了解,以避免不必要的麻烦。
总之,氘代DMSO作为一种重要的氘代溶剂,在化学位移研究中具有广泛的应用。
13CNMR核磁共振碳谱化学位移总览表

1) INEPT法
由于核磁共振本身信号灵敏度很低,尤其是低天然丰度的核 (如13C、15N等)更为突出。INEPT法是在具有两种核自旋的系统 中,以CH为例,通过脉冲技术,把高灵敏1H核的自旋极化传递到 低灵敏的13C核上去,这样由1H到与其偶合的13C的完全极化传递可 使,13C信号强度增强4倍。
的峰的裂分应全部去除。如果还有谱线的裂分不能去除,应考虑分
子中是否含F或P等元素。 (6)从分子式和可能的结构单元,推出可能的结构式。利用化学位移 规律和经验计算式,估算各碳的化学位移,与实测值比较。 (7)综合考虑1H NMR、IR、MS和UV的分析结果,必要时进行其他 的双共振技术及τ 1测定,排除不合理者,得到正确的结构式。
δ值范围在100-150ppm,sp杂化碳的δ值范围在60-95ppm。
2.诱导效应
当电负性大的元素或基团与碳相连时,诱导效应使碳的核外 电子云密度降低,故具有去屏蔽作用。随着取代基电负性增强, 或取代基数目增大,去屏蔽作用也增强, δ值愈向低场位移。
3.共轭效应
共轭作用会引起电子云分布的变化,导致不同位置碳的共 振吸收峰向高场或低场移动。
5.弛豫时间τ1可作为化合物结构鉴定的波谱参数
在化合物中,处于不同环境的13C核,它们的弛豫时间τ1数
值相差较大,可达2-3个数量级,通过τ1可以指认结构归属,
窥测体系运动状况等。
4.2.1
脉冲傅里叶变换法
原理同1H NMR。
4.2.2
核磁共振碳谱中几种去偶技术
在有机化合物的13C NMR中,13C-13C之间的偶合由于13C的天然丰 度很低,可以不予考虑。但13C-1H核之间的偶合常数很大,如1JCH高达 120-320Hz,13C的谱线会被与之偶合的氢按n+1规律裂分成多重峰,这 种峰的裂分对信号的归属是有用的,但当谱图复杂时,加上2JCCH、
核磁共振氢谱和碳谱

核磁共振氢谱核磁共振---NMR1945年美国斯坦福大学的 F. Block 和哈佛大学的 E. M. Purcell 同时发现了核磁共振现象,并因此荣获了1952年的 Nobel 物理奖。
核磁共振谱可为化合物鉴定提供下列信息:1.磁核的类型:由化学位移来判别,如在1HNMR 中,可判别甲基氢、芳氢、烯氢、醛氢等。
2.磁核的化学环境:由偶合常数和自旋-自旋裂分来判别,如在 1H-NMR 中可判定甲基是与-CH 2-相连,还是与苯环相连。
3.各类磁核的相对数量:氢谱中,通过积分面积或积分曲线来判断。
4 .核自旋弛豫时间:13CNMR 可提供 T 1,并用于结构归属指定,构象的测定,以及窥测体 系的运动情况。
5 .核间相对距离:通过核的 Overhause 效应可测得。
3.1核磁共振的基本原理3.1.1原子核的磁矩原子核是带正电荷的粒子,自旋将产生磁矩,但并非所有同位素的原子核有自旋,只有有自旋才有磁矩。
具有自旋运动的原子核具有一定自旋量子数(I ),I=1/2 *n ,那1,2,3···1. 核电荷数和和质量数均为偶数的原子核没自旋。
2. 核电荷数为奇数或偶数,核质量数为奇数,有自旋现象。
3. 核电荷数为奇数,核质量数为偶数,I 为整数的原子核有自旋现象。
对于自旋不为零的核来说,当其自旋时由于形成环电流,故而产生一个小磁场,这个小磁场可用核磁矩 μ 表示。
μ 是矢量,其大小由下式确定:πγγμ2)1(hI I p +==式中 γ ---核的磁旋比 p---自旋角动量不同的核有不同的 γ 值,是确定同位素核的特征常数。
3.1.2自旋核在磁场中的取向和能级对于I 不为零的核来说,如果不受外来磁场的干扰,其自旋轴的取向将是任意的。
当它们处于外加静磁场(磁场强度为H0)中时,根据量子力学理论,它们的自旋轴的取向不再是任意的,而只有(2I+1)种,这叫核自旋的空间量子化。
每一种取向可用一个磁量子数m 表示,则m=I,I-1,I-2,…-I+1,-I。
核磁氢谱中常见的官能团化学位移
核磁氢谱中常见的官能团化学位移核磁共振(Nuclear Magnetic Resonance,简称NMR)是一种重要的分析技术,可以用于研究化合物分子结构和化学环境。
在核磁共振谱中,氢原子的化学位移是一个重要的参数,可以提供分子中不同官能团的信息。
官能团是分子中具有特定化学性质的结构单元,每个官能团都有其特有的核磁化学位移范围。
下面介绍一些核磁氢谱中常见的官能团及其化学位移范围:1. 烷基(Alkyl)官能团烷基官能团是由碳和氢组成的烷烃分子链,例如甲基(CH3-)和乙基(C2H5-)等。
其化学位移范围通常在0.5~2.5 ppm之间。
2. 烯烃(Alkenyl)官能团烯烃官能团是含有碳—碳双键的分子,例如乙烯(C2H4-)和丙烯(C3H6-)等。
其化学位移范围通常在4.5~6.0 ppm之间。
3. 酮(Ketone)官能团酮官能团是由碳氧双键连接碳原子形成的结构,例如丙酮(CH3COCH3)和己酮(C5H9COCH3)等。
其化学位移范围通常在2.1~2.4 ppm之间。
4. 醇(Alcohol)官能团醇官能团是由羟基(-OH)连接到碳原子的结构,例如甲醇(CH3OH)和乙醇(C2H5OH)等。
其化学位移范围通常在0.5~5.0 ppm之间。
5. 醛(Aldehyde)官能团醛官能团是由碳氧双键和氢连接到同一个碳原子的结构,例如乙醛(CH3CHO)和丁醛(C4H9CHO)等。
其化学位移范围通常在9.0~10.0 ppm之间。
除了这些常见的官能团,核磁氢谱中还存在其他许多官能团化学位移,如羧酸、酰胺、卤代烷等。
通过对氢原子的化学位移的分析,我们可以进一步确定化合物的结构和化学环境。
CNMR核磁共振碳谱化学位移总览表课件
根据碳碳耦合和耦合常数,确定邻近碳原子 的类型和它们之间的关系。
峰形和强度
通过峰形和相对强度,确定化合物的结构和 它们在分子中的位置。
峰的位置和积分峰面积
通过峰的相对位置和积分峰面积,计算化合 物中各类碳原子的数量。
化学位移的基本概念和计算方法
1 化学位移
化学位移是一种定量指标,描述碳原子的磁场感受性,通常表示为δ值。
CNMR核磁共振碳谱化学 位移总览表课件
CNMR核磁共振碳谱化学位移总览表课件。
什么是核磁共振碳谱
核磁共振碳谱(CNMR)是一种分析化学的技术,用于确定和解释有机化合 物中碳原子的化学环境。
CNMR谱图的读取与解读方法
峰高与化学位移
测量CNMR谱图中峰的高度,根据化学位移 数值判断化合物中的碳原子类型。
2 计算方法
化学位移可以通过将每个峰位置相对于标准物质(如三甲基硅烷)进行测定和计算得到。
常见化合物的谱图和化学位移数值对照 表
谱图解读
通过解读常见化合物的CNMR谱图,学习化学位 移数值在不同类型化合物中的变化规律。
数值对照表
提供一张常见化合物的CNMR化学位移数值对照 表,帮助快速确定未知化合物的结构。
影响化学位移的因素及其解释
电子效应
电子的吸引或排斥会改变碳 原子的化学位移。
取代基效应
取代基的种类和位置会对化 学位移产生影响。
溶剂效应
溶剂分子的影响会改变化学 位移数值。
核磁共振碳谱在有机化学和药物研发中 的应用
1
结构确认
核磁共振碳谱帮助鉴组分定量
通过测量峰面积,核磁共振碳谱可以用于定量分析。
3
改进合成路径
通过核磁共振碳谱的结果,调整合成路径以获得更理想的产物。
13c nmr①碳谱的化学位移
O
O
中
C
学
99.98
O
O
C
1H
M
天然丰度/%
国
大
学
同位素
中
M
学
大
中
国
M
学
国
大
中
C
国
大
C
O
13C
M
O
大
学
国
O
C
O
大
学
M
国
中
O
O
C
M
国
大
学
中
O
中
O
C
O
大
学
M
国
C
O
M
O
大
学
国
中
学
学
M
M
M
大
学
大
大
国
国
国
中
中
中
M
M
M
学
学
学
大
大
大
国
国
国
中
中
中
M
M
学
学
大
大
国
国
中
中
碳谱的化学位移
1.核磁共振碳谱的特点
碳-13:丰度低,磁旋比小→灵敏度低
中
O
C
乙酸乙酯的1H/13C NMR图(溶剂CDCl3)
O
大
学
M
国
中
O
O
C
M
国
大
学
中
1H
国
C
O
M
O
大
学
国
中
溶剂的化学位移(碳谱和氢谱)
NMR Chemical Shifts of Common Laboratory Solvents as Trace Impurities Hugo E.Gottlieb,*Vadim Kotlyar,andAbraham Nudelman*Department of Chemistry,Bar-Ilan University,Ramat-Gan52900,IsraelReceived June27,1997In the course of the routine use of NMR as an aid for organic chemistry,a day-to-day problem is the identifica-tion of signals deriving from common contaminants (water,solvents,stabilizers,oils)in less-than-analyti-cally-pure samples.This data may be available in the literature,but the time involved in searching for it may be considerable.Another issue is the concentration dependence of chemical shifts(especially1H);results obtained two or three decades ago usually refer to much more concentrated samples,and run at lower magnetic fields,than today’s practice.We therefore decided to collect1H and13C chemical shifts of what are,in our experience,the most popular “extra peaks”in a variety of commonly used NMR solvents,in the hope that this will be of assistance to the practicing chemist.Experimental SectionNMR spectra were taken in a Bruker DPX-300instrument (300.1and75.5MHz for1H and13C,respectively).Unless otherwise indicated,all were run at room temperature(24(1°C).For the experiments in the last section of this paper,probe temperatures were measured with a calibrated Eurotherm840/T digital thermometer,connected to a thermocouple which was introduced into an NMR tube filled with mineral oil to ap-proximately the same level as a typical sample.At each temperature,the D2O samples were left to equilibrate for at least 10min before the data were collected.In order to avoid having to obtain hundreds of spectra,we prepared seven stock solutions containing approximately equal amounts of several of our entries,chosen in such a way as to prevent intermolecular interactions and possible ambiguities in assignment.Solution1:acetone,tert-butyl methyl ether,di-methylformamide,ethanol,toluene.Solution2:benzene,di-methyl sulfoxide,ethyl acetate,methanol.Solution3:acetic acid,chloroform,diethyl ether,2-propanol,tetrahydrofuran. Solution4:acetonitrile,dichloromethane,dioxane,n-hexane, HMPA.Solution5:1,2-dichloroethane,ethyl methyl ketone, n-pentane,pyridine.Solution6:tert-butyl alcohol,BHT,cyclo-hexane,1,2-dimethoxyethane,nitromethane,silicone grease, triethylamine.Solution7:diglyme,dimethylacetamide,ethyl-ene glycol,“grease”(engine oil).For D2O.Solution1:acetone, tert-butyl methyl ether,dimethylformamide,ethanol,2-propanol. Solution2:dimethyl sulfoxide,ethyl acetate,ethylene glycol, methanol.Solution3:acetonitrile,diglyme,dioxane,HMPA, pyridine.Solution4:1,2-dimethoxyethane,dimethylacetamide, ethyl methyl ketone,triethylamine.Solution5:acetic acid,tert-butyl alcohol,diethyl ether,tetrahydrofuran.In D2O and CD3OD nitromethane was run separately,as the protons exchanged with deuterium in presence of triethylamine.ResultsProton Spectra(Table1).A sample of0.6mL of the solvent,containing1µL of TMS,1was first run on its own.From this spectrum we determined the chemical shifts of the solvent residual peak2and the water peak. It should be noted that the latter is quite temperature-dependent(vide infra).Also,any potential hydrogen-bond acceptor will tend to shift the water signal down-field;this is particularly true for nonpolar solvents.In contrast,in e.g.DMSO the water is already strongly hydrogen-bonded to the solvent,and solutes have only a negligible effect on its chemical shift.This is also true for D2O;the chemical shift of the residual HDO is very temperature-dependent(vide infra)but,maybe counter-intuitively,remarkably solute(and pH)independent. We then added3µL of one of our stock solutions to the NMR tube.The chemical shifts were read and are presented in Table 1.Except where indicated,the coupling constants,and therefore the peak shapes,are essentially solvent-independent and are presented only once.For D2O as a solvent,the accepted reference peak(δ)0)is the methyl signal of the sodium salt of3-(trimeth-ylsilyl)propanesulfonic acid;one crystal of this was added to each NMR tube.This material has several disadvan-tages,however:it is not volatile,so it cannot be readily eliminated if the sample has to be recovered.In addition, unless one purchases it in the relatively expensive deuterated form,it adds three more signals to the spectrum(methylenes1,2,and3appear at2.91,1.76, and0.63ppm,respectively).We suggest that the re-sidual HDO peak be used as a secondary reference;we find that if the effects of temperature are taken into account(vide infra),this is very reproducible.For D2O, we used a different set of stock solutions,since many of the less polar substrates are not significantly water-soluble(see Table1).We also ran sodium acetate and sodium formate(chemical shifts: 1.90and8.44ppm, respectively).Carbon Spectra(Table2).To each tube,50µL of the stock solution and3µL of TMS1were added.The solvent chemical shifts3were obtained from the spectra containing the solutes,and the ranges of chemical shifts(1)For recommendations on the publication of NMR data,see: IUPAC Commission on Molecular Structure and Spectroscopy.Pure Appl.Chem.1972,29,627;1976,45,217.(2)I.e.,the signal of the proton for the isotopomer with one less deuterium than the perdeuterated material,e.g.,C H Cl3in CDCl3or C6D5H in C6D6.Except for CHCl3,the splitting due to J HD is typically observed(to a good approximation,it is1/6.5of the value of the corresponding J HH).For CHD2groups(deuterated acetone,DMSO, acetonitrile),this signal is a1:2:3:2:1quintet with a splitting of ca.2 Hz.(3)In contrast to what was said in note2,in the13C spectra the solvent signal is due to the perdeuterated isotopomer,and the one-bond couplings to deuterium are always observable(ca.20-30Hz). Figure1.Chemical shift of H DO as a function of tempera-ture..Chem.1997,62,7512-7515S0022-3263(97)01176-6CCC:$14.00©1997American Chemical Societyshow their degree of variability.Occasionally,in order to distinguish between peaks whose assignment was ambiguous,a further1-2µL of a specific substrate were added and the spectra run again.Table1.1H NMR Dataproton mult CDCl3(CD3)2CO(CD3)2SO C6D6CD3CN CD3OD D2O solvent residual peak7.26 2.05 2.507.16 1.94 3.31 4.79 H2O s 1.56 2.84a 3.33a0.40 2.13 4.87acetic acid CH3s 2.10 1.96 1.91 1.55 1.96 1.99 2.08 acetone CH3s 2.17 2.09 2.09 1.55 2.08 2.15 2.22 acetonitrile CH3s 2.10 2.05 2.07 1.55 1.96 2.03 2.06 benzene CH s7.367.367.377.157.377.33tert-butyl alcohol CH3s 1.28 1.18 1.11 1.05 1.16 1.40 1.24 OH c s 4.19 1.55 2.18tert-butyl methyl ether CCH3s 1.19 1.13 1.11 1.07 1.14 1.15 1.21 OCH3s 3.22 3.13 3.08 3.04 3.13 3.20 3.22 BHT b ArH s 6.98 6.96 6.877.05 6.97 6.92OH c s 5.01 6.65 4.79 5.20ArCH3s 2.27 2.22 2.18 2.24 2.22 2.21ArC(CH3)3s 1.43 1.41 1.36 1.38 1.39 1.40chloroform CH s7.268.028.32 6.157.587.90 cyclohexane CH2s 1.43 1.43 1.40 1.40 1.44 1.451,2-dichloroethane CH2s 3.73 3.87 3.90 2.90 3.81 3.78 dichloromethane CH2s 5.30 5.63 5.76 4.27 5.44 5.49diethyl ether CH3t,7 1.21 1.11 1.09 1.11 1.12 1.18 1.17 CH2q,7 3.48 3.41 3.38 3.26 3.42 3.49 3.56 diglyme CH2m 3.65 3.56 3.51 3.46 3.53 3.61 3.67 CH2m 3.57 3.47 3.38 3.34 3.45 3.58 3.61OCH3s 3.39 3.28 3.24 3.11 3.29 3.35 3.37 1,2-dimethoxyethane CH3s 3.40 3.28 3.24 3.12 3.28 3.35 3.37 CH2s 3.55 3.46 3.43 3.33 3.45 3.52 3.60 dimethylacetamide CH3CO s 2.09 1.97 1.96 1.60 1.97 2.07 2.08 NCH3s 3.02 3.00 2.94 2.57 2.96 3.31 3.06NCH3s 2.94 2.83 2.78 2.05 2.83 2.92 2.90 dimethylformamide CH s8.027.967.957.637.927.977.92 CH3s 2.96 2.94 2.89 2.36 2.89 2.99 3.01CH3s 2.88 2.78 2.73 1.86 2.77 2.86 2.85 dimethyl sulfoxide CH3s 2.62 2.52 2.54 1.68 2.50 2.65 2.71 dioxane CH2s 3.71 3.59 3.57 3.35 3.60 3.66 3.75 ethanol CH3t,7 1.25 1.12 1.060.96 1.12 1.19 1.17 CH2q,7d 3.72 3.57 3.44 3.34 3.54 3.60 3.65OH s c,d 1.32 3.39 4.63 2.47ethyl acetate CH3CO s 2.05 1.97 1.99 1.65 1.97 2.01 2.07C H2CH3q,7 4.12 4.05 4.03 3.89 4.06 4.09 4.14CH2C H3t,7 1.26 1.20 1.170.92 1.20 1.24 1.24 ethyl methyl ketone CH3CO s 2.14 2.07 2.07 1.58 2.06 2.12 2.19C H2CH3q,7 2.46 2.45 2.43 1.81 2.43 2.50 3.18CH2C H3t,7 1.060.960.910.850.96 1.01 1.26 ethylene glycol CH s e 3.76 3.28 3.34 3.41 3.51 3.59 3.65“grease”f CH3m0.860.870.920.860.88CH2br s 1.26 1.29 1.36 1.27 1.29n-hexane CH3t0.880.880.860.890.890.90CH2m 1.26 1.28 1.25 1.24 1.28 1.29HMPA g CH3d,9.5 2.65 2.59 2.53 2.40 2.57 2.64 2.61 methanol CH3s h 3.49 3.31 3.16 3.07 3.28 3.34 3.34 OH s c,h 1.09 3.12 4.01 2.16nitromethane CH3s 4.33 4.43 4.42 2.94 4.31 4.34 4.40 n-pentane CH3t,70.880.880.860.870.890.90CH2m 1.27 1.27 1.27 1.23 1.29 1.292-propanol CH3d,6 1.22 1.10 1.040.95 1.09 1.50 1.17 CH sep,6 4.04 3.90 3.78 3.67 3.87 3.92 4.02 pyridine CH(2)m8.628.588.588.538.578.538.52 CH(3)m7.297.357.39 6.667.337.447.45CH(4)m7.687.767.79 6.987.737.857.87 silicone grease i CH3s0.070.130.290.080.10 tetrahydrofuran CH2m 1.85 1.79 1.76 1.40 1.80 1.87 1.88 CH2O m 3.76 3.63 3.60 3.57 3.64 3.71 3.74 toluene CH3s 2.36 2.32 2.30 2.11 2.33 2.32CH(o/p)m7.177.1-7.27.187.027.1-7.37.16CH(m)m7.257.1-7.27.257.137.1-7.37.16 triethylamine CH3t,7 1.030.960.930.960.96 1.050.99 CH2q,7 2.53 2.45 2.43 2.40 2.45 2.58 2.57a In these solvents the intermolecular rate of exchange is slow enough that a peak due to HDO is usually also observed;it appears at2.81and3.30ppm in acetone and DMSO,respectively.In the former solvent,it is often seen as a1:1:1triplet,with2J H,D)1Hz. b2,6-Dimethyl-4-tert-butylphenol.c The signals from exchangeable protons were not always identified.d In some cases(see note a),the coupling interaction between the CH2and the OH protons may be observed(J)5Hz).e In CD3CN,the OH proton was seen as a multiplet atδ2.69,and extra coupling was also apparent on the methylene peak.f Long-chain,linear aliphatic hydrocarbons.Their solubility in DMSO was too low to give visible peaks.g Hexamethylphosphoramide.h In some cases(see notes a,d),the coupling interaction between the CH3and the OH protons may be observed(J)5.5Hz).i Poly(dimethylsiloxane).Its solubility in DMSO was too low to give visible peaks.Notes .Chem.,Vol.62,No.21,19977513.Chem.,Vol.62,No.21,1997NotesTable2.13C NMR Data aCDCl3(CD3)2CO(CD3)2SO C6D6CD3CN CD3OD D2O solvent signals77.16(0.0629.84(0.0139.52(0.06128.06(0.02 1.32(0.0249.00(0.01206.26(0.13118.26(0.02acetic acid CO175.99172.31171.93175.82173.21175.11177.21 CH320.8120.5120.9520.3720.7320.5621.03 acetone CO207.07205.87206.31204.43207.43209.67215.94 CH330.9230.6030.5630.1430.9130.6730.89 acetonitrile CN116.43117.60117.91116.02118.26118.06119.68 CH3 1.89 1.12 1.030.20 1.790.85 1.47 benzene CH128.37129.15128.30128.62129.32129.34tert-butyl alcohol C69.1568.1366.8868.1968.7469.4070.36 CH331.2530.7230.3830.4730.6830.9130.29 tert-butyl methyl ether OCH349.4549.3548.7049.1949.5249.6649.37 C72.8772.8172.0472.4073.1774.3275.62C C H326.9927.2426.7927.0927.2827.2226.60 BHT C(1)151.55152.51151.47152.05152.42152.85C(2)135.87138.19139.12136.08138.13139.09CH(3)125.55129.05127.97128.52129.61129.49C(4)128.27126.03124.85125.83126.38126.11CH3Ar21.2021.3120.9721.4021.2321.38C H3C30.3331.6131.2531.3431.5031.15C34.2535.0034.3334.3535.0535.36chloroform CH77.3679.1979.1677.7979.1779.44cyclohexane CH226.9427.5126.3327.2327.6327.961,2-dichloroethane CH243.5045.2545.0243.5945.5445.11 dichloromethane CH253.5254.9554.8453.4655.3254.78diethyl ether CH315.2015.7815.1215.4615.6315.4614.77 CH265.9166.1262.0565.9466.3266.8866.42 diglyme CH359.0158.7757.9858.6658.9059.0658.67 CH270.5171.0369.5470.8770.9971.3370.05CH271.9072.6371.2572.3572.6372.9271.63 1,2-dimethoxyethane CH359.0858.4558.0158.6858.8959.0658.67 CH271.8472.4717.0772.2172.4772.7271.49 dimethylacetamide CH321.5321.5121.2921.1621.7621.3221.09 CO171.07170.61169.54169.95171.31173.32174.57NCH335.2834.8937.3834.6735.1735.5035.03NCH338.1337.9234.4237.0338.2638.4338.76 dimethylformamide CH162.62162.79162.29162.13163.31164.73165.53 CH336.5036.1535.7335.2536.5736.8937.54CH331.4531.0330.7330.7231.3231.6132.03 dimethyl sulfoxide CH340.7641.2340.4540.0341.3140.4539.39 dioxane CH267.1467.6066.3667.1667.7268.1167.19 ethanol CH318.4118.8918.5118.7218.8018.4017.47 CH258.2857.7256.0757.8657.9658.2658.05 ethyl acetate C H3CO21.0420.8320.6820.5621.1620.8821.15 CO171.36170.96170.31170.44171.68172.89175.26CH260.4960.5659.7460.2160.9861.5062.32CH314.1914.5014.4014.1914.5414.4913.92 ethyl methyl ketone C H3CO29.4929.3029.2628.5629.6029.3929.49 CO209.56208.30208.72206.55209.88212.16218.43C H2CH336.8936.7535.8336.3637.0937.3437.27CH2C H37.868.037.617.918.148.097.87 ethylene glycol CH263.7964.2662.7664.3464.2264.3063.17“grease”CH229.7630.7329.2030.2130.8631.29n-hexane CH314.1414.3413.8814.3214.4314.45CH2(2)22.7023.2822.0523.0423.4023.68CH2(3)31.6432.3030.9531.9632.3632.73HMPA b CH336.8737.0436.4236.8837.1037.0036.46 methanol CH350.4149.7748.5949.9749.9049.8649.50c nitromethane CH362.5063.2163.2861.1663.6663.0863.22 n-pentane CH314.0814.2913.2814.2514.3714.39CH2(2)22.3822.9821.7022.7223.0823.38CH2(3)34.1634.8333.4834.4534.8935.302-propanol CH325.1425.6725.4325.1825.5525.2724.38 CH64.5063.8564.9264.2364.3064.7164.88 pyridine CH(2)149.90150.67149.58150.27150.76150.07149.18 CH(3)123.75124.57123.84123.58127.76125.53125.12CH(4)135.96136.56136.05135.28136.89138.35138.27 silicone grease CH3 1.04 1.40 1.38 2.10 tetrahydrofuran CH225.6226.1525.1425.7226.2726.4825.67 CH2O67.9768.0767.0367.8068.3368.8368.68 toluene CH321.4621.4620.9921.1021.5021.50C(i)137.89138.48137.35137.91138.90138.85CH(o)129.07129.76128.88129.33129.94129.91CH(m)128.26129.03128.18128.56129.23129.20CH(p)125.33126.12125.29125.68126.28126.29triethylamine CH311.6112.4911.7412.3512.3811.099.07 CH246.2547.0745.7446.7747.1046.9647.19a See footnotes for Table1.b2J PC)3Hz.c Reference material;see text.For D2O solutions there is no accepted reference for carbon chemical shifts.We suggest the addition of a drop of methanol,and the position of its signal to be defined as49.50ppm;on this basis,the entries in Table2were recorded.The chemical shifts thus obtained are,on the whole,very similar to those for the other solvents. Alternatively,we suggest the use of dioxane when the methanol peak is expected to fall in a crowded area of the spectrum.We also report the chemical shifts of sodium formate(171.67ppm),sodium acetate(182.02and 23.97ppm),sodium carbonate(168.88ppm),sodium bicarbonate(161.08ppm),and sodium3-(trimethylsilyl)-propanesulfonate[54.90,19.66,15.56(methylenes1,2, and3,respectively),and-2.04ppm(methyls)],in D2O. Temperature Dependence of HDO Chemical Shifts.We recorded the1H spectrum of a sample of D2O, containing a crystal of sodium3-(trimethylsilyl)propane-sulfonate as reference,as a function of temperature.The data are shown in Figure1.The solid line connecting the experimental points corresponds to the equation which reproduces the measured values to better than1 ppb.For the0-50o C range,the simplergives values correct to10ppb.For both equations,T is the temperature in°C.Acknowledgment.Generous support for this work by the Minerva Foundation and the Otto Mayerhoff Center for the Study of Drug-Receptor Interactions at Bar-Ilan University is gratefully acknowledged.JO971176Vδ)5.060-0.0122T+(2.11×10⁻⁵)T2(1)δ)5.051-0.0111T(2)Notes .Chem.,Vol.62,No.21,19977515。